Direct arylations of indoles and pyrroles with differently substituted diaryliodonium salts were shown to efficiently proceed in the absence of metal catalysts.
A new Pd-catalyzed reaction for the coupling between perfluoroalkyl iodides (RFI) and simple aromatic substrates is described. The perfluoroalkylated arene products are obtained in good to excellent yields in the presence of a phosphine-ligated Pd catalyst and Cs2CO3 as a base. The development, optimization, scope, and preliminary mechanistic studies of these transformations are reported.
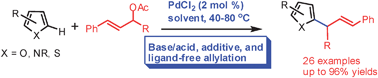
The Author present a PdCl2-catalyzed protocol for highly efficient allylation and benzylation of a rich variety of N-, O-, and S-containing heteroarenes under base/acid, additive, and ligand-free conditions. The method represents the very few examples for simple, universally applicable, clean, and atom-efficient functionalization of heteroarenes.
Nickeleophilic addition! The first example of a nickel-catalyzed direct intramolecular nucleophilic addition of aryl or vinyl chlorides to α-ketoamides through CCl bond activation is reported, which takes place under mild reaction conditions (see scheme).
Ketones and alkynes join together: The catalytic title reaction proceeds rapidly at 120 °C and requires only 1 hour for completion (see scheme). The reaction mechanism likely involves chelation-assisted ortho CH activation, insertion of the alkyne moiety, carbocyclization, and protonation. Cp*=pentamethylcyclopentadienyl.
It appears that transition metal catalysts are not necessary to perform the direct arylation of electron-rich heterocycles with aryl iodides and bromides. Lithium tert-butoxide in DMF promotes this reaction for a variety of N-alkyl- and N-arylpyrroles as well as for benzofuran and some other electron-rich aromatic compounds and provides the desired products in moderate to high yields. In contrast to all previous reports on the Pd-catalyzed direct arylation of indolizine, the reaction mediated by lithium tert-butoxide proceeds selectively at position 5.
In the absence of transition-metal catalysts, a Mizoroki–Heck-type reaction proceeded to give stilbene derivatives in a simple manner using an aryl halide, a styrene derivative, KOtBu, EtOH, and DMF (see scheme; DMF=N,N-dimethylformamide).
Removable and versatile: The 2-pyridylsulfinyl group has proved to be an efficient directing group in the PdII-catalyzed aryl ortho CH olefination. This catalyst system enables the sequential double olefination to give asymmetrically di-ortho-functionalized arenes. The sulfinyl directing group can be easily cleaved, providing access to 1,3-disubstituted arenes, or transformed into a thiol group.
This paper presents a detailed investigation of the factors controlling site selectivity in the Pd-mediated oxidative coupling of 1,3-disubstituted and 1,2,3-trisubstituted arenes (aryl−H) with cyclometalating substrates (LC−H). The influence of both the concentration and the steric/electronic properties of the quinone promoter are studied in detail. In addition, the effect of steric/electronic modulation of the carboxylate ligand is discussed. Finally, we demonstrate that substitution of the carboxylate for a carbonate X-type ligand leads to a complete reversal in site selectivity for many arene substrates. The origins of these trends in site selectivity are discussed in the context of the mechanism of Pd-catalyzed oxidative cross-coupling.
A transition-metal-free method for arylation of heterocycle and arene carbon−hydrogen bonds by aryl chlorides and fluorides has been developed. The reactions proceed via aryne intermediates and are highly regioselective with respect to the C−H bond coupling component.
A new catalytic protocol for the regioselective direct arylation of quinoline derivatives at the 8-position has been developed. The reaction is catalyzed by a Rh(NHC) system, and the choice of the NHC ligand was most important for achieving high reactivity and selectivity.